Your structure viewer shows a gap between two well-resolved segments. A preparation tool offers to rebuild it. Should you accept the repair before docking, mutation design, or simulation?
Model a missing segment when the downstream question requires coordinates there and you can represent the uncertainty honestly. Do not fill every gap merely to make the structure look complete. A plausible loop is a computational hypothesis, not a recovered experimental observation.
Start by establishing what is missing. An unresolved segment, an engineered deletion, absent side-chain atoms, and a break caused by file handling are different problems. They need different responses.
Key takeaways
- Establish whether the gap represents unresolved coordinates, a real sequence deletion, or a file-processing artifact.
- Rebuild only when the question requires it, and preserve the provenance of added coordinates.
- Test whether conclusions survive plausible alternatives; good geometry does not prove experimental occupancy.
Missing coordinates do not necessarily mean missing sequence
Experimental structures do not always include coordinates for every residue in the sample. A flexible or heterogeneous region may lack sufficient evidence for one supported conformation. Conversely, the experimental construct may genuinely omit a segment found in the full biological sequence.
The RCSB guide to missing coordinates explains this distinction. Consult the deposited polymer sequence, construct information, and relevant annotations alongside the visible model.
Also inspect numbering carefully. A jump from residue 100 to 110 might indicate an unresolved stretch, an intentional deletion, or a numbering convention. Counting missing integers is not a reliable reconstruction of the experimental sequence.
Check whether the apparent gap arose during export. Chain selection, removal of nonstandard residues, or conversion between file formats can change what your downstream software sees. Preserve the original downloaded structure so you can distinguish deposited information from preparation artifacts.
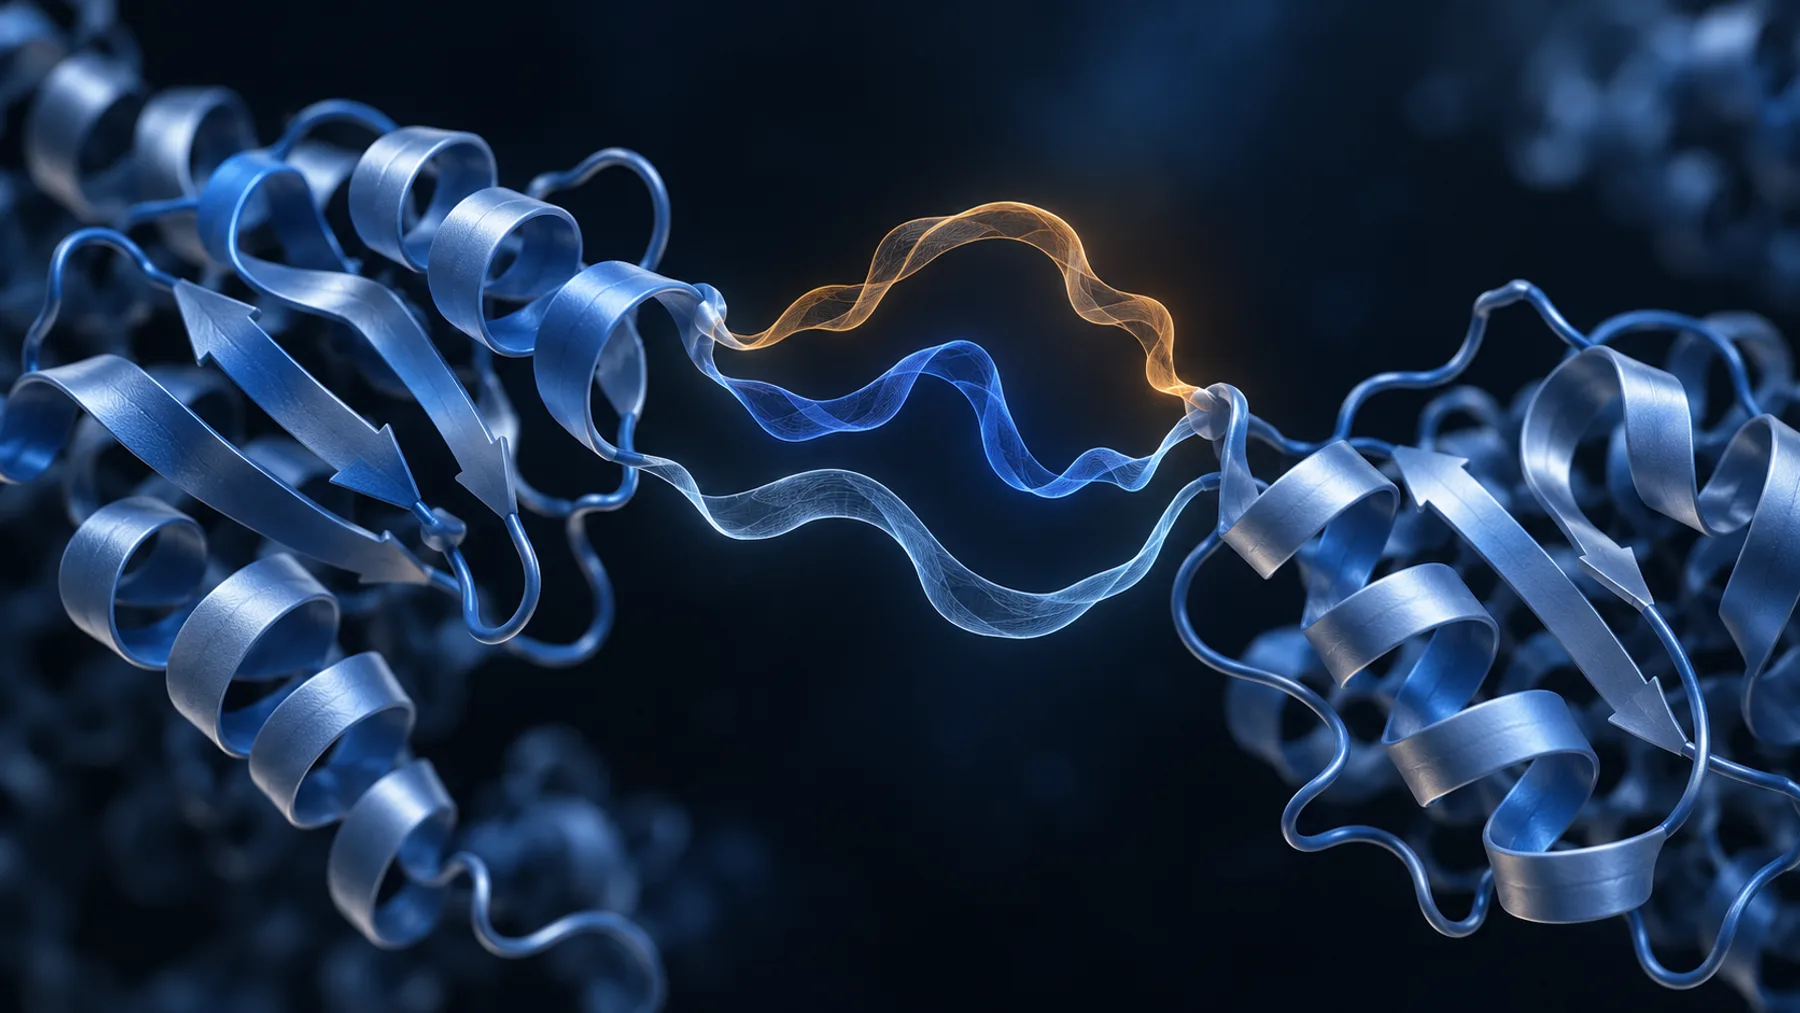
Figure 1. An unresolved segment is different from a sequence deletion. Faint paths indicate uncertainty, not observed coordinates. Conceptual illustration.
Classify the gap before choosing a modeling method
| Situation | Appropriate first action |
|---|---|
| Present in the construct sequence but unresolved | Assess whether its uncertainty affects the planned analysis |
| Absent from the experimental construct | Decide whether you are intentionally modeling a different construct |
| Missing side-chain atoms only | Rebuild the required atoms with suitable geometry and inspect the local environment |
| File conversion or chain-selection artifact | Correct the preparation or import step before modeling |
| Uncertain mapping to the biological sequence | Resolve sequence and residue identifiers first |
The RCSB primary-sequence guide helps reconcile sequence records with coordinate content. The wwPDB annotation documentation provides further detail on deposited information and annotation.
For an internal gap, identify both flanking residues, their geometry, and the missing sequence. For a terminal segment, recognize that there may be fewer geometric constraints. A long terminal tail can have many plausible conformations even when the neighboring domain is well resolved.
What the experimental absence does and does not tell you
Missing coordinates are an absence of a supported atomic model for a region in that experiment. They can be consistent with flexibility or heterogeneity, but they do not uniquely identify intrinsic disorder. Experimental resolution, local occupancy, sample composition, and model-building choices can also matter. Treat the missing segment as a question to investigate, not as a complete biological annotation.
An unresolved loop and a high-confidence predicted loop may therefore coexist without either record being internally inconsistent. The experimental structure describes what was supported in a particular sample and state. The prediction proposes a conformation under its learned assumptions. A prediction can be useful without explaining why the experimental region was unresolved.
Inspect evidence at the level relevant to the claim. A deposited model and its validation report help identify what was modeled and how well supported the surrounding region is. Where available and appropriate, the underlying density or map can reveal whether there is local signal that is incompatible with a single clean conformation. Absence of coordinates alone cannot answer that question.
The RCSB missing-coordinate guide is useful because it separates the polymer sequence from the residues represented in the coordinate model. Carry that distinction into every derived file. If a preparation tool inserts coordinates, the sequence may become continuous, but the added region has not acquired experimental support merely by being written beside observed atoms.
For a long terminal segment, the relevant uncertainty may be an ensemble rather than one missing loop. Drawing one compact tail can imply a fixed contact that the evidence does not establish. If the project only requires the resolved domain, restricting the claim to that domain may be more defensible than constructing a detailed but weakly constrained tail.
Ask whether the gap can change the conclusion
If you are inspecting a buried mutation far from an unresolved terminal tail, rebuilding the tail may add little useful information. You can often preserve the observed domain and state the limitation.
If the missing loop forms a pocket entrance, contacts a ligand, participates in an interface, or lies near a proposed mutation, its uncertainty may determine the result. Omitting it could create an artificial cavity or expose a surface that is normally covered. Filling it with one arbitrary conformation can create a different artifact.
For a hypothetical docking project, a missing surface loop on the opposite side of the protein may be low priority after checking spatial context. A missing loop that borders the docking box deserves explicit alternative hypotheses or further evidence before a large screen.
Our structure-based drug-design guide explains why receptor preparation must follow the biological question rather than a generic cleanup checklist.
Choose the strongest available source of constraints
Look for another experimental structure of the same target, a relevant state, or a close homologue with a comparable segment. Check sequence differences, construct boundaries, ligands, crystal contacts, and domain arrangements before transferring coordinates.
A homologue's resolved loop is useful evidence, but it may represent a different state or be stabilized by contacts absent from your system. Likewise, a predicted full-length model may supply a plausible hypothesis without establishing that the loop adopts that conformation experimentally.
Inspect prediction confidence and the relative placement of the modeled segment. Our explanation of pLDDT and PAE is relevant here. Low confidence should remain visible in the downstream interpretation; it should not disappear because the segment was pasted into an otherwise experimental structure.
If evidence is weak, generate or retain a small set of physically plausible alternatives rather than presenting one as definitive. The alternatives should represent meaningful uncertainty, not merely duplicate one solution with tiny coordinate perturbations.
Match the modeling approach to the constraint you actually have
There are several ways to propose missing coordinates. A related experimental structure can supply a candidate conformation; a loop-modeling method can search conformations consistent with the flanking structure; a full-protein predictor can suggest a broader arrangement. The choice should follow the source of the uncertainty, rather than whichever tool offers the fastest “repair” button.
The loop-modeling work of Fiser, Do and Sali (2000) explicitly evaluated how prediction accuracy depends on factors including loop length and the correctness of the surrounding structure. The lesson for interpretation is that fixed endpoints and a low score do not provide uniform accuracy across all gaps. A short, constrained segment and a long loop beside an uncertain domain arrangement are different problems.
For template transfer, compare the local context before accepting the coordinates. A loop held in place by a ligand, a binding partner, or crystal packing may not represent the intended state. Sequence similarity can support the relationship while leaving the conformation unresolved. Explain which constraint the template contributes and which assumptions remain.
For a predicted full-length model, first reconcile sequence and domain placement with the experimental structure. Superimposing one domain can leave another displaced, making a visually plausible patch geometrically inconsistent at its boundaries. The experimental and predicted models may also represent different conformational states. A merged file should not silently hide that disagreement.
For de novo loop alternatives, diversity is useful only if the candidates remain physically and biologically plausible. A collection of severe clashes is not an informative uncertainty ensemble. At the other extreme, many near-identical outputs from one starting assumption do not establish that other plausible conformations were explored. Describe the scope of the alternatives and avoid presenting their frequency as a calibrated probability of occupancy.
Validate geometry and contacts after rebuilding
Check chain connectivity, bond geometry, stereochemistry, clashes, and compatibility with neighboring atoms. Inspect whether the loop makes implausible contacts with ligands, membranes, symmetry-related molecules, or other chains relevant to the analysis.
Preserve the experimental portion deliberately. A repair procedure that extensively rearranges well-supported regions can change the original question. Record any relaxation or minimization and inspect the important changes.
Passing geometric checks means the model is physically more plausible. It does not prove that the conformation is occupied in the experiment. Several different loops can satisfy the same endpoints and avoid clashes.
Mark rebuilt residues in the saved model or accompanying metadata. Keep the original structure, the missing-segment sequence, the modeling method, software version, settings, and model identifiers. A colleague should be able to separate observed coordinates from added hypotheses without comparing files atom by atom.
For simulation, completeness still requires a biological decision
Molecular simulation often requires a chemically defined system with suitable connectivity and termini. That requirement does not tell you which unresolved loop conformation is correct.
Depending on the scientific question, you may model the full segment, use a justified truncated construct with appropriate terminal treatment, or compare alternative initial conformations. Each choice changes the simulated system and must be stated.
Do not assume that a short equilibration will erase uncertainty in an arbitrarily rebuilt loop. Relevant rearrangements may not occur on the simulated timescale. If the reported result depends strongly on the initial loop choice, that dependence belongs in the conclusion.
Test sensitivity before trusting the repaired model
Repeat the key analysis across the relevant alternatives when feasible. Compare pocket access, contacts, mutation environments, or other outputs that matter to the project. Keep all other settings consistent so the effect of the missing segment is interpretable.
If the decision is stable across reasonable alternatives, you have evidence that the gap is not dominating that particular conclusion. If the decision changes, report the uncertainty and identify the experiment or additional structure that would resolve it.
For example, a proposed mutation may remain solvent exposed in every plausible loop model. That is a more robust interpretation than a contact observed in only one arbitrarily rebuilt loop. Conversely, a ligand pose that exists only when the loop is omitted needs a clear qualification.
Figure 2. Alternative loop placements can change the interpretation of a nearby pocket. Testing sensitivity helps reveal whether the missing segment drives the conclusion. Conceptual illustration.
Worked example: a loop beside a binding pocket
Imagine a hypothetical experimental structure with an unresolved segment near a pocket entrance. The target sequence and deposited polymer record agree that the segment was present. A related structure provides one possible closed conformation, while an independently generated model suggests a more open placement. Both can be made geometrically plausible, but neither is directly observed in the sample of interest.
| Structural interpretation | What it can test | Limitation to retain |
|---|---|---|
| Original structure with the gap | Whether the observed pocket core supports the proposed interaction | The omitted segment may expose an artificial opening |
| Plausible closed-loop model | Whether the proposed interaction survives a more restricted entrance | The closed state is a hypothesis for this sample |
| Plausible open-loop model | Whether an alternative placement changes pocket access or contacts | Geometric plausibility does not establish occupancy |
Suppose an interaction with the resolved pocket core is retained in both loop alternatives, but a proposed contact with the loop occurs in only one. Those conclusions have different support. The core interaction is less sensitive to the sampled loop uncertainty; the loop-specific contact remains conditional. Neither finding proves binding, but the comparison prevents a single repaired model from carrying more weight than it deserves.
If the whole proposed binding mode exists only when the loop is omitted, the gap dominates the interpretation. A larger docking calculation using the same incomplete receptor would repeat the assumption at greater scale. Additional structural evidence or a different experimental question may be more useful than generating more scores.
Before interpreting those differences, inspect the local geometry. Chen et al. (2010) describe MolProbity's use of all-atom contacts and stereochemical criteria for structure validation. Such checks help reject implausible models. They do not establish which of several acceptable loop conformations belongs to the biological ensemble.
Keep the original and derived coordinates separately, and make the uncertainty visible in the accompanying figure or table. A practical handoff states the gap, its sequence mapping, the evidence used to propose alternatives, the validation performed, and which conclusions changed. That record lets the next researcher decide whether the model is adequate for their question without inheriting an unexplained repaired structure.
Keep a repair record with the design
Orbion Characterize can support structural review, and Orbion Design can help organize candidate designs. Keep added coordinates and their provenance visible in the evidence used to choose experiments. A visually complete structure should not imply uniformly strong evidence across every residue.
The useful endpoint is a model appropriate to the question, with known limitations. Sometimes that means a carefully rebuilt loop. Sometimes it means leaving a gap, restricting the analysis, or postponing a conclusion until the missing region is better constrained.
Common questions
Should I always replace a missing loop with AlphaFold coordinates?
No. First verify sequence mapping, state, local confidence, and compatibility with the experimental structure. Predicted coordinates remain predictions after they are transferred into an experimental model.
Is a low-energy loop the correct loop?
Not necessarily. An energy or scoring function evaluates a model under its assumptions. It does not establish experimental occupancy, and several plausible conformations may remain.
Can I leave the gap for docking?
Only after assessing whether it affects the pocket, preparation requirements, or receptor chemistry. Document the omission and test sensitivity when the segment could influence the result.
References
- RCSB PDB-101. Missing coordinates. Official guide. Accessed October 2026.
- RCSB PDB-101. Primary sequences. Official guide. Accessed October 2026.
- wwPDB. Biocuration procedures and policies. Annotation documentation. November 2022 version.
- Fiser A, Do RK, Sali A. Modeling of loops in protein structures. Protein Science. 2000. doi:10.1110/ps.9.9.1753
- Chen VB, et al. MolProbity: all-atom structure validation for macromolecular crystallography. Acta Crystallographica D. 2010. doi:10.1107/S0907444909042073